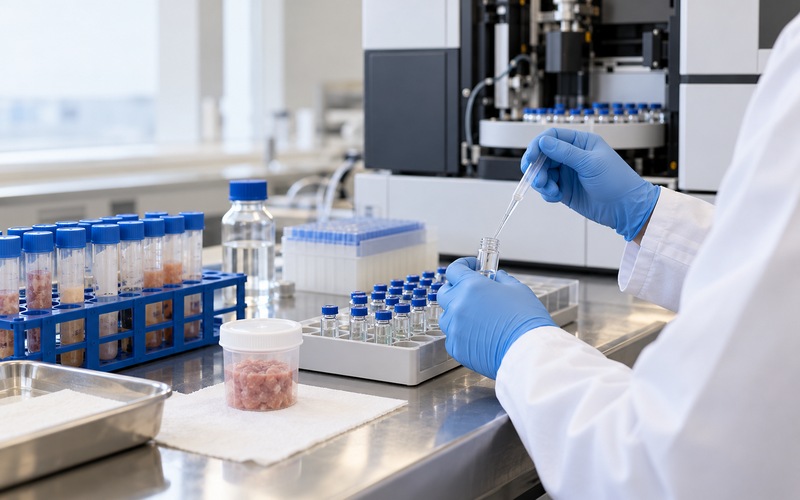
A defensible veterinary drug residue result begins before the vial reaches the autosampler. Laboratories need a scope tied to the correct species, tissue, analyte, and decision level, then a confirmation workflow that controls extraction, matrix effects, identification evidence, calibration, and the samples affected by any failure.
Key Points
Liquid chromatography with tandem mass spectrometry, or LC-MS/MS, is well suited to multi-residue work because it can combine chromatographic separation with selective mass transitions. That capability can tempt a laboratory to treat one extraction and one acquisition method as a universal answer. Veterinary drug residue testing resists that shortcut. A cattle kidney sample, a swine muscle sample, and a milk sample can present different recovery, interference, and regulatory questions even when the target compound is the same.
The current U.S. Department of Agriculture Food Safety and Inspection Service Chemistry Laboratory Guidebook lists CLG-MRM3, Screening and Confirmation of Animal Drug Residues by UHPLC-MS/MS, with a September 1, 2026, effective date. FSIS announced that the 2026 revision extended screening and confirmation to ovine, caprine, and porcine muscle and kidney, following earlier additions for bovine matrices. That method is a valuable regulatory reference. It is not automatic validation for a private laboratory, another jurisdiction, an unlisted species, or a different matrix.
Start With a Five-Part Decision Framework
Identify the jurisdiction, commodity program, species, production class, edible tissue, marker residue, and action or reporting level.
State whether the method screens, confirms, quantifies, or performs more than one function. List every matrix and analyte included in the claim.
Consider water, fat, protein, endogenous compounds, drug metabolites, tissue heterogeneity, storage, and expected concentration.
Specify calibration, identification criteria, blanks, spikes, internal standards, reference materials, carryover checks, and continuing verification.
Document how presumptive results, confirmed results, inconclusive results, QC failures, reruns, and amended reports will be handled.
The first step prevents one of the costliest errors in residue testing: building the method around an instrument panel rather than the legal and scientific decision. Part 556 defines a tolerance as the maximum concentration of a marker residue, or another named monitoring residue, that may legally remain in a specified edible tissue. The same regulation distinguishes marker residue, target tissue, edible tissue, and total residue. Those are operational definitions. They determine what should be measured and what inference can be made from the result.
For example, a tolerance linked to a target tissue may support an inference about other edible tissues under the regulation. When a tolerance is not expressly linked to a target tissue, Part 556 says that inference cannot be made. A laboratory information management system should therefore preserve species, tissue, analyte, and decision basis as structured data rather than reduce the sample to a generic “animal tissue” label.
Decide What Screening and Confirmation Must Each Accomplish
| Workflow Element | Primary Purpose | Operational Advantage | Main Risk to Control |
|---|---|---|---|
| Broad multi-residue screen | Detect a large panel efficiently and identify samples that need further work. | High coverage and faster triage across many analyte classes. | A presumptive signal may lack the selectivity or performance evidence required for a final decision. |
| Targeted LC-MS/MS confirmation | Establish identity and, when required, concentration using predefined chromatographic and mass-spectral evidence. | Focused review of transitions, ion relationships, retention behavior, calibration, and controls. | Confirmation criteria can fail even when a peak appears at the expected time. |
| Quantitative confirmation | Compare a confirmed residue with the applicable tolerance, action level, or program rule. | Produces a decision-ready result when the measurand and calibration are correctly aligned. | Bias from extraction, matrix effects, calibration, or an incorrect marker residue can change the disposition. |
| Orthogonal or alternate confirmation | Resolve an interference or strengthen identity evidence when the primary procedure is inconclusive. | Can separate a genuine residue from a method-specific artifact. | An alternate result is only useful when the second procedure is qualified for the stated purpose. |
The laboratory should set the handoff between these stages in advance. A screen can be designed for sensitivity and coverage, but a presumptive response should not quietly become a reportable confirmation because the chromatogram looks convincing. Confirmation needs its own acceptance criteria, calibrators, quality controls, integration review, and sequence logic.
For laboratories supporting European Union work, Commission Implementing Regulation (EU) 2021/808 provides binding performance and interpretation requirements for analytical methods used for residues of pharmacologically active substances. Its identification criteria, validation design, decision concepts, and quality-control expectations belong to that regulatory context. They can inform method development elsewhere, but a laboratory should not present them as U.S. requirements or combine criteria from different frameworks without a documented scientific and contractual basis.
Choose Sample Preparation Around the Matrix and Analyte Set
Extraction is where method scope becomes physical. Protein precipitation may be efficient for a broad panel, while liquid-liquid extraction can improve cleanup for selected chemistries. Solid-phase extraction may provide greater control over selectivity and concentration, but it adds sorbent behavior, conditioning, loading, wash, and elution steps. Dilute-and-shoot approaches reduce transfers and cost, yet they can send more matrix into the LC-MS/MS system and increase suppression, contamination, and maintenance.
| Preparation Strategy | Best Fit | Tradeoff | Evidence to Request |
|---|---|---|---|
| Protein precipitation and centrifugation | Broad polar-to-moderate panels where throughput matters and the extract remains manageable. | Fast, but residual phospholipids and coextractives may affect ionization and source cleanliness. | Matrix-effect profiles, recovery by analyte class, extract stability, and maintenance trend data. |
| Liquid-liquid extraction | Analytes that partition predictably under controlled pH and solvent conditions. | Useful cleanup, with risks from emulsions, phase error, evaporation, and variable recovery across chemistries. | pH robustness, phase-volume control, recovery across matrices, and reconstitution performance. |
| Solid-phase extraction | Targeted panels requiring stronger cleanup or concentration. | More selective and potentially cleaner, but more steps and susceptibility to cartridge lot or loading effects. | Breakthrough, capacity, lot verification, recovery, wash selectivity, and automation compatibility. |
| QuEChERS-style partitioning or dispersive cleanup | Multi-residue workflows where partitioning and rapid cleanup can cover diverse compounds. | Broad and efficient, though pH, salts, hydration, sorbent selection, and analyte stability need control. | Analyte-class recovery, salt and sorbent lot checks, matrix effects, and extract compatibility. |
| Dilution after a simple extraction | Robust, sensitive assays where sensitivity can be exchanged for lower matrix load. | Reduces matrix effects and contamination, but may move low-level residues below the verified reporting capability. | Dilution integrity, detection capability at the decision level, and internal-standard behavior. |
Method developers should challenge the preparation with representative matrix variation, not only a single clean lot. Kidney and liver may differ from muscle in protein, lipid, endogenous metabolites, and background interferences. Milk and eggs introduce their own composition and homogenization questions. Where blank matrix is scarce, document the source, suitability, and limitations of the material used for validation and calibration.
Build Calibration and Matrix Controls Together
Calibration answers how response relates to amount under defined conditions. It does not prove that the sample was homogeneous, the residue was recovered, or the matrix did not alter ionization. Those risks require separate controls. The FDA’s Foods Program Guidelines for the Validation of Chemical Methods provide a useful primary framework for fit-for-purpose validation in food, feed, cosmetics, and veterinary products. Laboratories should use the current controlling edition and their program requirements when setting acceptance criteria.
A resilient quantitative design may combine isotopically labeled internal standards, matrix-matched calibration or a justified alternative, fortified matrix controls, independent check standards, method blanks, and dilution checks. The mix depends on the method claim. An internal standard can correct some variability, but only when it is appropriate for the analyte and added early enough to experience the relevant losses. It cannot compensate reliably for every extraction failure, interference, or analyte-specific transformation.
Matrix effects should be evaluated as a distribution across representative lots and matrices. A mean near 100% can hide severe suppression in one lot and enhancement in another. Examine analyte response, internal-standard response, recovery before and after extraction, and the residual effect after the planned dilution. If matrix-matched calibrators are used, establish that the blank matrix is appropriate and does not contain the analyte or an interfering signal.
Define Confirmation Evidence Before Reviewing Peaks
A confirmation decision should combine chromatographic and mass-spectral evidence. The procedure should state the required retention-time agreement, monitored transitions or accurate-mass ions, relative ion response, signal-to-noise or other detection evidence, integration rules, and how interferences are recognized. The exact criteria belong to the governing method and program. They should not be invented during review or relaxed because a result is expected.
Analysts also need a clear inconclusive path. A sample may show a response at the expected retention time while failing an ion-ratio criterion, containing an unresolved interference, or exceeding the validated range. “Detected” is not automatically “confirmed,” and “not confirmed” is not automatically “absent.” The final wording should reflect what the method demonstrated and what it did not.
Design the Sequence to Reveal Contamination and Drift
- Verify system readiness. Confirm maintenance state, column condition, source cleanliness, tune or suitability status, mobile phases, and sequence configuration.
- Establish the calibration. Review individual levels, fit, residuals, back-calculated performance, internal-standard response, and any predefined exclusion rules.
- Demonstrate cleanliness. Place solvent and method blanks where they can reveal preparation contamination and instrument memory.
- Challenge the batch. Include fortified controls, independent checks, duplicates or replicate preparations, and reference materials according to the validated plan.
- Bracket reportable samples. Use continuing checks and strategically placed blanks so the effect of drift or carryover can be bounded.
- Close the review. Reconcile injections, reinjections, integrations, sequence changes, audit-trail events, and sample disposition before release.
Carryover deserves specific attention because residue panels can span a wide concentration range. A high residue or fortified control can contaminate the needle, seat, tubing, injector, or column. A blank after a high sample helps reveal that risk, but the laboratory also needs a response plan. That plan should identify which following samples are held, what cleaning and verification are required, and when reinjection is scientifically meaningful.
Common Failure Modes and the First Useful Question
| Observed Failure | Likely Risk Areas | First Useful Question |
|---|---|---|
| Fortified matrix control recovers low | Extraction loss, analyte instability, adsorption, incorrect spike timing, matrix suppression. | Did the internal standard fail in the same direction, and was it added before the step suspected of causing loss? |
| Internal-standard response is low only in certain samples | Sample-specific suppression, preparation error, injection problem, or interference. | Does dilution restore response, and is the affected sample matrix within the validated claim? |
| Qualifier relationship fails | Interference, low signal, transition error, saturation, integration, or wrong analyte identity. | Do the raw traces support identity under the approved criteria, or must the result remain inconclusive? |
| Blank after a high sample contains analyte | Autosampler or column carryover, preparation contamination, or solvent contamination. | Which subsequent samples lack a clean blank boundary and must be held? |
| Continuing check drifts during the batch | Source contamination, calibration solution change, mobile phase, column condition, temperature, or detector response. | When did the shift begin, and which samples are bracketed by acceptable controls? |
| Duplicate preparations disagree | Heterogeneous tissue, subsampling, weighing, incomplete homogenization, extraction, or dilution. | Do replicate injections agree while independent preparations diverge? |
| Result is above the calibration range | Unexpected residue, inadequate dilution design, or detector saturation. | Is a validated dilution available, and does it preserve the confirmation evidence and decision level? |
The first investigation should preserve the original evidence. Review the raw data, sequence, preparation record, instrument state, maintenance history, standards, and audit trail before choosing a repeat. Repeated injections until a preferred answer appears can erase the distinction between a diagnostic experiment and result selection. The quality system should specify who authorizes additional work and how the original and repeat data are linked.
Illustrative Example: Ceftiofur in Bovine Kidney
Hypothetical planning example. A laboratory is validating a targeted LC-MS/MS confirmation for a ceftiofur marker residue in bovine kidney. The team begins with the current U.S. tolerance and the controlling program method, then defines the marker residue, target tissue, reporting units, decision level, and confirmation criteria.
Kidney lots are assessed for endogenous interference and matrix-effect variability. The preparation includes a fortified matrix control, a separately prepared check, an appropriate internal standard added before extraction, a method blank, and a dilution check. The sequence places a blank after the highest calibrator and after a high fortified control.
During a routine batch, the fortified control meets the recovery rule, but the blank after a high sample contains a confirmed analyte signal. The laboratory holds the following samples that lack a clean boundary, documents the carryover investigation, cleans and verifies the injection path, and repeats only the affected work under the approved procedure. It does not change the confirmation criteria or discard the original data.
This is an editorial example, not a regulatory method or a universal acceptance scheme. The current regulation, program method, validated laboratory procedure, and contract determine the actual criteria and disposition.
Actionable Implementation Checklist
- Record the jurisdiction, program, species, production class, tissue, analyte, marker residue, and decision level.
- State separately what the method screens, confirms, and quantifies.
- Verify the current official method revision before transferring or modifying a published procedure.
- Define representative matrix lots, blank-matrix suitability, storage, homogenization, and test-portion controls.
- Compare preparation options using recovery, matrix effects, selectivity, throughput, contamination risk, and maintenance burden.
- Justify internal-standard selection and addition timing for each analyte class.
- Validate calibration range, model, weighting, dilution integrity, and decision-level capability.
- Write confirmation criteria and the inconclusive-result path before routine testing.
- Place blanks and continuing checks so drift and carryover have a clear sample boundary.
- Predefine control failures, holds, diagnostic experiments, rerun authority, and report amendment.
- Preserve raw data, original integrations, sequence changes, audit trails, and the link between original and repeat work.
- Review changes in official methods, tolerances, analyte panels, species coverage, and matrix claims through change control.
Related LabPress Reading
For instrument-selection questions, see Best LC-MS Systems for Regulated Quality-Control Laboratories. For a closer look at extraction tradeoffs and matrix effects, read Choosing Sample Preparation Methods for Environmental LC-MS. The matrix is different, but the decision discipline around dilution, solid-phase extraction, and suppression remains useful.
Sources
- USDA FSIS Chemistry Laboratory Guidebook, including current CLG-MRM3 listing.
- USDA FSIS Constituent Update, August 28, 2026, describing the September 2026 matrix expansion.
- FDA Foods Program Guidelines for the Validation of Chemical Methods in Food, Feed, Cosmetics, and Veterinary Products.
- FDA Foods Program Compendium of Analytical Laboratory Methods.
- 21 CFR Part 556, Tolerances for Residues of New Animal Drugs in Food.
- Commission Implementing Regulation (EU) 2021/808, consolidated version.
What Would You Change First?
Where does your veterinary residue workflow carry the greatest unresolved risk: matrix coverage, extraction recovery, ion suppression, confirmation evidence, carryover, or the investigation path after a failed control?
Keep Exploring
Related Reading
Choosing Sample Preparation Methods for Environmental LC-MS
Compare dilution, SPE, and liquid-liquid extraction for environmental LC-MS, with practical checks for recovery, matrix effects, and throughput.Read Article
Designing a Cannabis Potency Testing Calibration and QC Strategy
A practical framework for cannabis potency calibration, reference standards, matrix-aware quality control, failure investigations, and defensible reporting.Read Article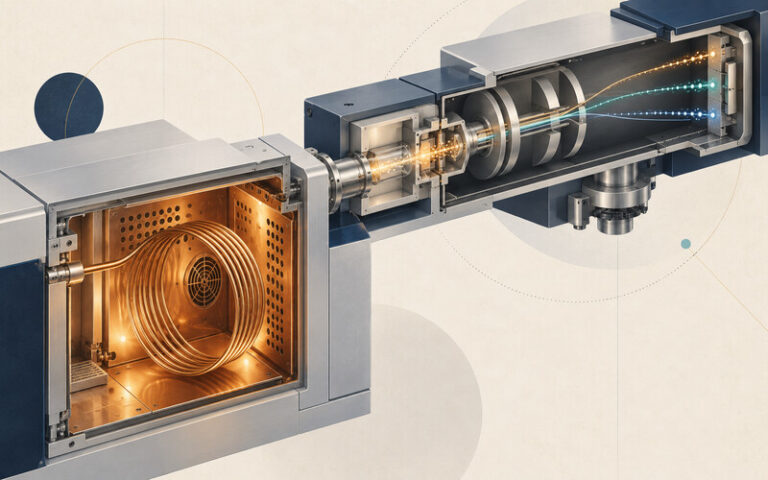
Leave a Reply